Focus keyword: fda 510k vs de novo hematology analyzer
Theme: Regulation & Compliance
Every US market analyst will tell you it is a simple choice between 510(k) and De Novo—until they meet a device that fits neither cleanly.
The 510(k) pathway works when a blood analyzer is substantially equivalent to an already-cleared predicate: similar technology, similar performance, similar intended use. De Novo exists for genuine low-to-moderate risk innovations with no predicate at all. The trap is that marketing a novel 5-part hematology analyzer as 510(k) with a stretched predicate can earn a refuse-to-accept letter, while filing De Novo when a predicate clearly exists wastes a year. Know your device before you pick a lane.
When 510(k) is the right call
A 5-part differential analyzer that uses established impedance plus optical scatter, with performance comparable to a predicate device, is a textbook 510(k) candidate. You need comparative data, bench studies and a truthful substantial-equivalence argument. This is the fastest and cheapest route, and it is the default for most incremental analyzer improvements.
When De Novo is unavoidable
If your analyzer introduces a new technology—say AI-assisted blast flagging that changes the clinical decision frame—there may be no predicate. De Novo lets you establish a new class, but it demands more clinical evidence and a longer review clock. The payoff is a clear regulatory path that no competitor can copy without their own De Novo.
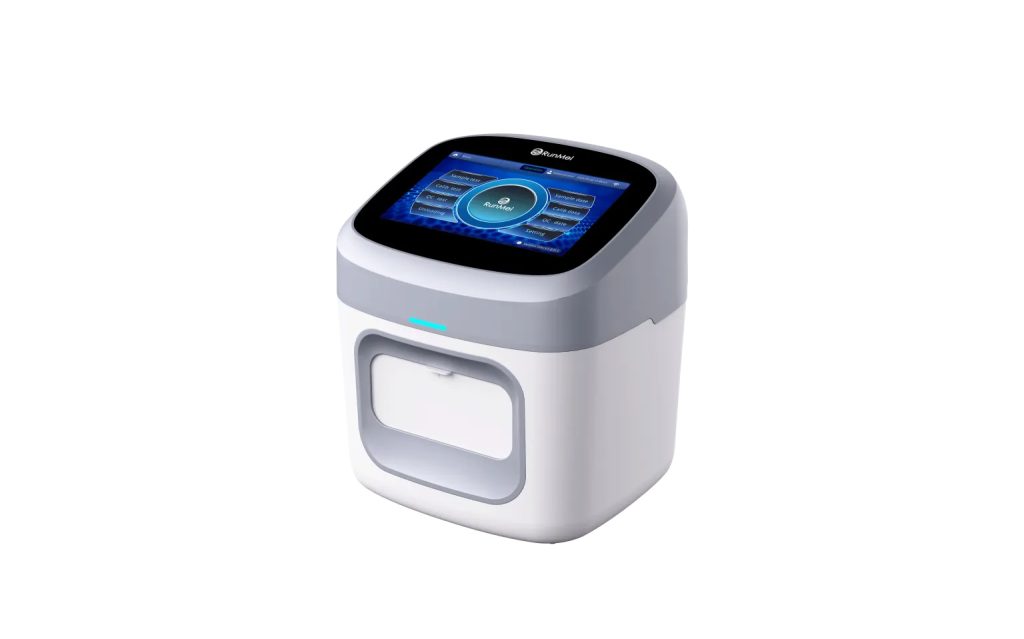
For a practical example in this space, see the RunMei diagnostic products.
Explore the RunMei Health.
As a next step, review the OEM/ODM manufacturing partner.
For reference, see the analyzer frequently asked questions.
Why This Matters: Authoritative Context
Regulatory context is grounded in primary sources: the EU In Vitro Diagnostic Regulation 2017/746 (IVDR) defines the transition and classification duties the industry tracks, the U.S. FDA’s 510(k) and De Novo pathways govern US market access under 21 CFR Part 820, and ISO 15189 and ISO 13485 set the quality and laboratory-management benchmarks referenced throughout. For WHO Prequalification, the WHO In Vitro Diagnostics and Laboratory Technology programme publishes the specifications and performance requirements that prequalification assessments apply. Consulting these primary texts keeps a sourcing or validation decision honest and defensible.